1. Executive Summary & Diagnostic Outsourcing Market Dynamics #
The global in vitro diagnostics (IVD) industry is undergoing a monumental paradigm shift. Driven by post-pandemic decentralized healthcare demands, expanding point-of-care (POCT) clinical testing, and the stringent regulatory barriers imposed by the European Union In Vitro Diagnostic Medical Devices Regulation (EU IVDR 2017/746) and FDA 21 CFR Part 820, diagnostic innovators increasingly rely on Contract Development and Manufacturing Organizations (CDMOs).
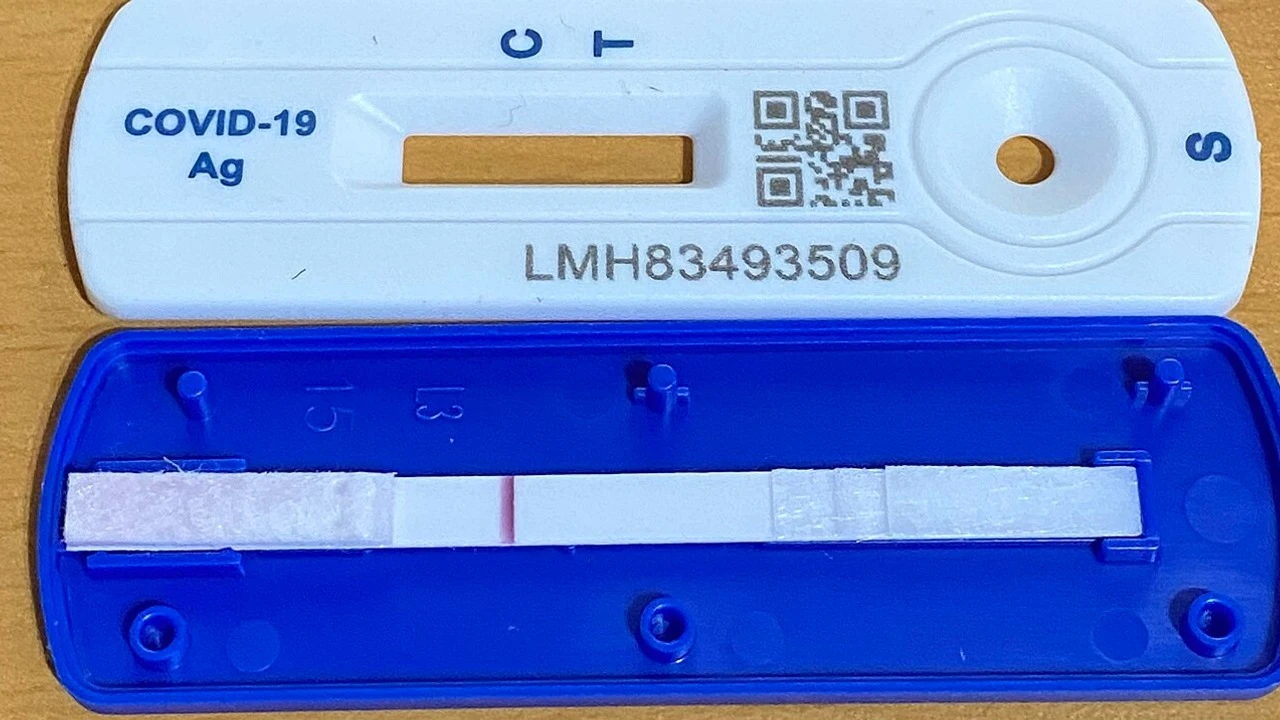
CDMO Audit Note — Rahul Kumar, Chief Editorial Director: "During a CDMO site audit, always ask to inspect the rejection bin next to the automated cassette press-closure station. If plastic housing pins are crushing the sample pad by even 0.25 mm too tight, capillary flow stalls mid-strip—creating a 15% invalid-rate nightmare in clinical field lots."
While diagnostic biotechs and molecular startups excel at novel biomarker discovery, antibody pairing, and proof-of-concept prototype fabrication, transitioning from manual benchtop assembly (producing 500 hand-cut strips per week) to commercial high-volume manufacturing (500,000 to 5,000,000 cassette tests per month) is fraught with commercial risk. Over 40% of diagnostic commercialization delays are directly attributable to failed technology transfers, inadequate cleanroom climate control, robotic dispensing variation, and incomplete Design History Files (DHF).
Selecting a qualified diagnostic CDMO is not a routine procurement decision; it is a long-term strategic partnership that determines product regulatory clearance, gross margins, and analytical consistency. This industrial guide provides diagnostic founders, chief technical officers (CTOs), and quality directors with an exhaustive engineering audit framework. We examine automated reel-to-reel continuous dispensing, laser guillotine cutting metrology, dry-room environmental physics (<10% RH), automated ultrasonic cassette swaging, and detailed ISO 13485:2016 audit checklists.
2. Core Manufacturing Technologies: Manual Assembly vs. Automated Continuous Reel-to-Reel #
The first fundamental technical discriminator when auditing an IVD CDMO is their level of physical process automation.
Diagnostic Manufacturing Automation Continuum:
[Tier 1: Manual / Batch] ──► [Tier 2: Semi-Automated Sheet] ──► [Tier 3: Continuous Reel-to-Reel]
 - Output: <50k tests/mo      - Output: 200k - 500k tests/mo      - Output: 1M - >10M tests/mo
 - Hand assembly              - Batch strip cutters               - Continuous tension control
 - High human error (CV>15%)  - Moderate labor (CV: 8-12%)        - Vision inspection (CV < 4%)
2.1 Limitations of Batch-Based Sheet Processing (Tier 1 & 2) #
In entry-level manufacturing, diagnostic strips are produced using precut plastic-backed cards (30 cm × 7.5 cm):
- Membranes, sample pads, conjugate pads, and absorbent wicks are laminated manually or semi-automatically by human operators aligning edges against mechanical fixtures.
- Reagents are striped across the 30 cm cards in discrete batches.
- Inherent Process Defects:
- Operator-dependent alignment error: A 0.5 mm variance in overlap between the conjugate pad and nitrocellulose membrane alters fluid capillary transfer by >30%, creating slow flow or sample pooling.
- End-of-strip edge artifacts: Start-and-stop dispense cycles generate uneven pressure at sheet borders, forcing the disposal of the first and last 5 strips per card (10% - 15% direct material scrap).
2.2 Continuous Reel-to-Reel Automated Manufacturing (Tier 3) #
World-class diagnostic CDMOs operate continuous web-handling lines (e.g., Kinematic Automation, BioDot, Kinematic Matrix lines):
- Raw materials (nitrocellulose rolls, backing cards, sample matrices) are fed continuously from large reels under closed-loop electronic tension control (0.5 N ± 0.05 N).
- Non-contact bio-jet or micro-solenoid dispensers deliver capture antibodies and colloidal conjugates onto the moving web at web speeds of 50 - 150 mm/sec with volumetric precision within ± 1.0%.
- Continuous inline vision inspection systems (high-speed line-scan CCD cameras) examine line width, line density, and laminate overlap tolerances in real time, automatically marking defective zones for pneumatic ejection downstream.
- Outcome: Production scale exceeding 100,000 finished tests per shift per machine line with analytical coefficients of variation (CV%) below 4.0%.
3. Cleanroom Environmental Physics: Dry-Room Engineering & Humidity Control #
The single most critical environmental factor governing lateral flow assay stability and coefficient of variation is relative humidity (RH).
Dry-Room Environmental Control Architecture:
Ambient Air (25°C, 60% RH)
       │
       ▼ (Chilled Glycol Pre-Cooling Coil: Dehumidifies to 10°C)
Dual-Bed Desiccant Rotor (Lithium Chloride / Molecular Sieve Matrix)
       │
       ▼ (Bake-Out Reactivation Zone Expels Moisture to Atmosphere)
Ultra-Dry Process Air (<10% RH, Dew Point: -40°C)
       │
       ▼ (HEPA H14 Filtration Unit)
Cleanroom Processing Suite (Positive Pressure: +25 Pa, 20°C ± 1°C, RH < 10%)
3.1 The Destructive Thermodynamics of Moisture on Nitrocellulose #
Nitrocellulose is an inherently hygroscopic, porous ester polymer. If nitrocellulose membranes or conjugate pads are exposed to ambient room humidity (>40% RH) during strip assembly:
- Water vapor condenses into the microscopic 8 - 15 mum pores, interacting with the surfactant wetting agents (e.g., Triton X-100) embedded in the membrane matrix.
- Capillary migration speed shifts unpredictably, altering the fluid residence time (tau = (L2 eta / 2 gamma r costheta)) of sample over the Test line.
- Dry lyophilized colloidal gold or fluorescent Europium conjugates prematurely re-hydrate, causing irreversible antibody denaturation and aggregation.
3.2 Audit Criteria for CDMO Environmental Control #
When touring a prospective CDMO facility, the audit team must physically inspect the environmental monitoring logs:
- Lamination & Assembly Suite: Must operate at ≤ 20% RH at 20°C ± 2°C.
- Conjugate Drying & Packaging Suite (Dry Room): Must maintain ≤ 10% RH (Dew Point ≤ -20°C) under active desiccant wheel dehumidification.
- Continuous Data Logging: Calibration records of multi-point capacitive hygrometers with automated alarm triggers must be documented in compliance with 21 CFR Part 11.
4. Key Automated Manufacturing Equipment to Audit #
During an on-site technical audit, examine the pedigree, age, and maintenance logs of five core automated subsystems:
| Equipment Category | Industry Standard Platforms | Critical Metrology Specifications | Failure Risk if Uncontrolled |
|---|---|---|---|
| Non-Contact Reagent Dispensers | BioDot BioJet Elite / XYZ3060; Kinematic Matrix | Volumetric accuracy ± 1.0%; line position tolerance ± 0.05 mm | Uneven line thickness; wandering test line resulting in automated optical reader errors |
| Continuous Reel-to-Reel Laminators | Kinematic Automation HR4000; BioDot Reel-to-Reel | Web tension control ± 0.05 N; laminate overlap accuracy ± 0.1 mm | Variable capillary flow, sample pooling, or total failure of fluid transfer between pads |
| High-Speed Rotary Guillotine Cutters | Kinematic 1200 / BioDot CM4000 | Strip width tolerance: 4.00 mm ± 0.05 mm; zero shear pinching | Crushed membrane edges causing rapid edge-wicking and distorted parabolic flow fronts |
| Automated Cassette Clamshell Assembly | Hauni / Harro Höfliger / Custom Vision Robotics | Press force monitoring (150 N ± 10 N); ultrasonic weld depth ± 0.02 mm | Crushed internal nitrocellulose or loose fit causing fluid bypass outside strip cavity |
| High-Speed Pouching & Packaging | Multivac / Bosch Packaging Systems | Hermetic heat-seal width ≥ 8 mm; inline desiccant pouch verification | Leaking pouch seals admitting moisture; loss of on-shelf stability (<24 months) |
5. Technology Transfer Engineering: The Stage-Gate Process #
A disciplined CDMO enforces a formal, phased Stage-Gate technology transfer framework compliant with GAMP 5 and ISO 13485:2016 Clause 7.3 (Design Transfer).
Stage-Gate Technology Transfer Roadmap:
[Gate 1: Feasibility / Analytical Verification] ──► Confirm formulation & pairing on pilot equipment
                                                                 │
                                                                 ▼
[Gate 2: Design for Manufacturing (DFM)] ──► Lock raw material BOM, tooling & tolerances
                                                                 │
                                                                 ▼
[Gate 3: Process Validation (IQ / OQ / PQ)] ──► Execute 3 consecutive commercial verification lots
                                                                 │
                                                                 ▼
[Gate 4: Commercial Scale cGMP Production] ──► Release for routine global clinical supply
5.1 Installation, Operational, and Performance Qualification (IQ/OQ/PQ) #
Before initiating full-scale manufacturing, teams must resolve assay chemistry sensitivities. When evaluating test line background yields, consult our bench protocol for troubleshooting high background noise and false positives in lateral flow assays. Furthermore, ensuring reagent conjugation stability requires raw materials utilizing carrier-free recombinant antigens for optimal conjugation efficiency and strict ISO 8655 pipette calibration compliance standards.
Before commercial manufacturing release, the CDMO must validate every automated production line:
- Installation Qualification (IQ): Verifies that equipment is installed according to engineering drawings, with correct electrical supplies, HEPA cleanroom exhausts, and pneumatic air filters (0.01 mum oil-free).
- Operational Qualification (OQ): Demonstrates that dispensing pumps, cutters, and vision systems operate within specified limits across upper and lower boundary conditions (e.g., testing strip cutting at 3.90 mm and 4.10 mm width; testing dispense rates at ± 15% nominal pressure).
- Performance Qualification (PQ): Execution of three (3) consecutive full-scale manufacturing lots using commercial raw materials. All lots must demonstrate:
- Process Capability Index: Cpk ≥ 1.33 for all critical quality parameters (line position, signal intensity, flow rate).
- Diagnostic Sensitivity: ≥ 98.0% against international clinical reference panels.
- Overall Manufacturing Yield: ≥ 95.0% (scrap rate ≤ 5.0%).
6. Comprehensive CDMO Audit Checklist & Scoring Framework #
Utilize this structured scoring matrix when conducting quality audits of candidate manufacturing facilities.
| Audit Domain | Essential Evaluation Criteria | Maximum Points | Passing Score |
|---|---|---|---|
| 1. Quality Management System (QMS) | ISO 13485:2016 certification by a recognized Notified Body (TÜV, BSI, DNV); FDA 21 CFR Part 820 registration; documented CAPA system with robust root-cause methodology | 25 | 22 |
| 2. Cleanroom Environmental Controls | Validated ISO Class 7/8 cleanrooms; continuous environmental monitoring (<10% RH dry rooms); automated HVAC alarm interlocks; HEPA air change frequency ≥ 20/hr | 20 | 18 |
| 3. Automated Production Hardware | Continuous reel-to-reel dispensing and lamination capability; automated inline vision systems; automated cassette press/welding lines with force transducers | 20 | 17 |
| 4. Raw Material Supply Chain & Traceability | Documented incoming raw material release testing (NC membrane porosity, backing card adhesive thickness, conjugate gold sphericity); secondary supplier qualification | 15 | 13 |
| 5. Design History File (DHF) & Tech Transfer | Robust Design Transfer procedures (Clause 7.3.8); clear IQ/OQ/PQ validation protocols; comprehensive batch records and automated electronic signatures (21 CFR Part 11) | 20 | 18 |
| Total Evaluation Score | Exhaustive Facility Audit | 100 | 88 |
Policy: Any finding classified as a "Critical Non-Conformity" (e.g., absence of dry-room humidity logs, unvalidated automated cutters, or uncalibrated pipetting systems) triggers an immediate audit failure regardless of aggregate numerical score.
Lateral Flow CDMO Quality Issues: Critical Failure Modes in Point of Care Diagnostics Manufacturing #
During technical transfer and commercial scale-up with a point of care diagnostics CDMO, lateral flow assays (LFAs) frequently exhibit catastrophic yield drops. Over 80% of post-transfer lot failures stem from four pervasive engineering and environmental quality issues:
1. Nitrocellulose Capillary Flow Rate Drift & Bedding Variations #
Master rolls of nitrocellulose membrane exhibit batch-to-batch capillary flow time variances (± 15% seconds per 4 cm). If the CDMO does not calibrate reel-to-reel tension and dispense heights dynamically, buffer front velocities skew across the master card, leading to incomplete conjugate mobilization, false negatives, or variable line intensities.
2. High-Throughput Biodot Dispensing CV% and Conjugate Aggregation #
Industrial reel-to-reel jet dispensing requires strict viscosity management. Colloidal gold and fluorescent latex microsphere conjugates subjected to excessive pneumatic pressure or uncalibrated piezoelectric nozzles experience shear-induced agglomeration. A qualified CDMO must demonstrate in-line optical CV < 3.5% across 100 meters of continuous dispensing.
3. Strip Guillotine Edge Fraying & Lateral Fluid Bypass #
Blunt cutting blades or improper blade clearance during automated rotary slitting tear the nitrocellulose backing and wick boundaries. Micro-fraying along the strip perimeter creates capillary bypass channels where sample fluid races along strip edges ahead of the membrane front, producing high background smearing and invalid test runs.
4. Foil Pouch Sealing Micro-Pores & Cleanroom Humidity Degradation #
Nitrocellulose and immobilized antibodies undergo rapid thermodynamic denaturing when exposed to relative humidity exceeding 20% RH. High-speed pouch sealing stations must undergo helium leak detection and dye ingress testing. CDMO dry-room airlocks must strictly maintain < 10% RH at 20°C during assembly, desiccant insertion, and foil pouch heat-sealing.
7. Comprehensive Troubleshooting: Resolving Common Post-Transfer Manufacturing Defects #
| Manufacturing Failure Mode | Underlying Root Cause | Diagnostic Evidence | Mandatory CDMO Corrective Action |
|---|---|---|---|
| Parabolic or "Crescent" fluid front during test run | Guillotine cutter blades worn or misaligned, creating crushed membrane edge fibers | Microscopic inspection of strip edge shows ragged fibers; flow travels faster along edges | Replace carbide cutting blades; verify blade clearance (<5 mum); realign strip guidance rollers |
| Sample pools at sample pad and fails to cross into membrane | Inadequate compression force between sample pad and nitrocellulose laminate | Fluid pauses at interface; no capillary migration down strip | Increase adhesive backing lamination pressure; increase pad overlap tolerance from 1.5 mm to 2.0 mm |
| Lot-to-lot baseline shift in automated optical reader values | Dispense pump volume drifting due to fluctuating supply web tension | Spectrophotometer confirms antibody line width varies along the reel | Implement closed-loop laser micrometer line-width tracking; calibrate web tension servo controllers |
| Rapid degradation of test line signal after 30 days accelerated stability | Moisture leakage through defective cassette blister or pouch heat-seal | Residual moisture in pouch >2.0 mg; desiccant pouch fully saturated | Validate pouch sealing temperature and pressure; increase desiccant mass from 0.5 g to 1.0 g silica gel |
| High cassette rejection rate due to membrane crushing | Clamshell plastic tooling shrinkage or excessive ultrasonic welding horn energy | Depressed indentations visible across nitrocellulose membrane upon opening cassette | Re-tool plastic injection molds with tighter shrinkage tolerances (± 0.05 mm); install physical mechanical depth stops |
8. Regulatory Compliance and International Standards #
- International Organization for Standardization (ISO). ISO 13485:2016: Medical devices — Quality management systems — Requirements for regulatory purposes.
- Food and Drug Administration (FDA). 21 CFR Part 820: Quality System Regulation for Medical Devices.
- European Union. Regulation (EU) 2017/746 on in vitro diagnostic medical devices (IVDR).
- International Society for Pharmaceutical Engineering (ISPE). GAMP 5: A Risk-Based Approach to Compliant GxP Computerized Systems.